Products
Sherpa
Drafter
Reviewer
Researcher
Audiences
MedTech
Startups
Consultants
Guideways.ai
About us
Technology
FAQ
Trust
News & Insights
Contact us

☰
An AI regulatory affairs agent is a specialised AI system that carries out defined regulatory and quality compliance tasks for medical devices. It can break a regulatory question into several steps, search relevant sources, apply task-specific logic and produce an evidence-backed output for review.
Medical device teams can use these agents for classification, pathway identification, regulatory research, document drafting, submission-readiness checks and quality management system gap analysis.
Their value depends on the quality of the regulatory logic, source material, traceability and review process. The medical device manufacturer remains responsible for regulatory decisions, evidence and documentation.
This article explains how AI regulatory affairs agents differ from generic chatbots and conventional software, why medical device teams are adopting them, which workflows they can support, how to assess their credibility, where human oversight remains essential, and how they fit within MedTech companies and regulatory consultancies.
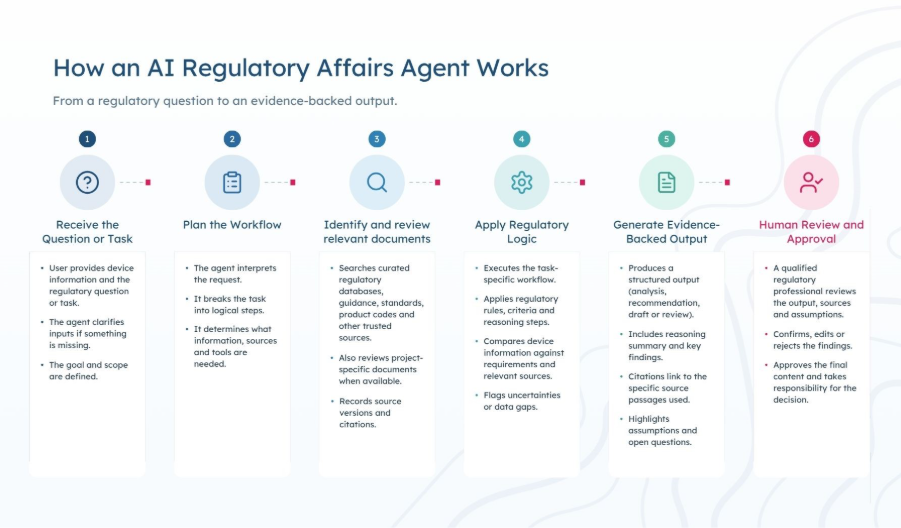
An AI regulatory affairs agent performs specialised regulatory work through a sequence of connected actions.
When a user submits a question, the agent can determine what information is needed, divide the task into stages, search relevant databases or documents and combine the findings into a structured output.
A specialised regulatory agent adds regulatory knowledge, task-specific workflows and access to appropriate reference material.
For example, when assessing whether a device fits an existing US product code, the analysis to consider:
A general AI assistant may identify broad similarities between devices. A regulatory agent should assess whether those similarities are relevant to the regulatory decision.
“Agent” describes how a system performs work. It does not confirm that its conclusions are correct.
A regulatory agent can still produce an incomplete or unsuitable output when:
Teams should assess the system’s reasoning, evidence and controls rather than rely on the product label.
Medical device regulation involves a connected set of decisions, requirements and evidence.
Before development progresses too far, a manufacturer needs to determine:
Every design iteration, risk management update, or post-market surveillance finding triggers a cascade that impacts several documents and new supporting evidence.
A revised intended purpose may alter classification. A newly identified hazard may require an additional risk control. That control may create a new design requirement, verification activity, usability consideration or clinical-evidence need.
Correcting one gap can therefore require updates across several documents and new supporting evidence.
Left unmanaged, this compounding administrative burden creates severe operational bottlenecks. As compliance requirements scale faster than internal teams can hire, manual document management leads to prolonged submission cycles, delayed market entry, and heightened risk of non-compliance errors.
Regulatory professionals and consultants work across large volumes of material while supporting several products, markets and internal stakeholders.
Manual research, document comparison and consistency review take time that experts could spend on interpretation, risk assessment and strategic decisions.
AI regulatory affairs agents can support teams by:
These benefits depend on the quality of the system and the review process around it.
Regulatory strategy affects product claims, development choices, evidence plans, budgets and timelines.
Starting regulatory analysis after the device has already been built can expose requirements that would have been easier to address during concept and design.
An AI regulatory affairs agent can help teams examine regulatory implications earlier, before product decisions become difficult or expensive to change.
General-purpose AI can support document summarisation, restructuring and language improvement. These tasks may not require a specialised regulatory system.
The limits become more significant when the tool is used to support a regulatory determination.
Regulatory analysis often depends on a defined sequence of questions.
Consider two software devices that analyse the same type of medical image. One outlines an area for a clinician to inspect. Another interprets the image and recommends a diagnosis.
The technologies may be related, but the clinical function, risk profile and evidence requirements differ.
A regulatory analysis needs to consider:
Broad similarity is not enough to support a regulatory conclusion.
Although most large language model have encountered regulatory documents during training, they do not retain every requirement as an exact and current reference.
A model can produce an answer that appears plausible while misstating a requirement, using outdated information or combining details from different sources.
For regulatory work, the system should retrieve the relevant current document, identify the applicable passage and show how that information informed the conclusion.
This is especially important when the task involves:
A team should be able to see:
Without this information, the team may need to repeat the analysis to verify the result.
The strongest use cases are structured, evidence-intensive tasks that can be divided into clear regulatory steps.
During the concept or early development stage, an agent may help organise and assess:
The output should include assumptions, unresolved questions and the basis for the proposed direction.
It should be reviewed by a qualified person and should not be treated as a binding decision from a regulator or notified body.
Supporting guide: AI Agent for Medical Device Classification and Regulatory Pathway Planning
Regulatory teams often need to answer questions spread across regulations, guidance, databases and internal documentation.
A research agent can:
Its value comes from searching within a defined regulatory context and relating the findings to a device or task.
Specialised agents can help create structured first drafts from information already established about the device.
Examples include:
A clinical evaluation plan depends on several upstream inputs, including:
A useful drafting workflow should build from those inputs rather than generate a generic template.
The manufacturer must review and own the content. Proposed risk controls, evidence plans and procedures affect real development and operational activities.
An AI agent may compare a developing submission with the requirements associated with the relevant FDA pathway.
Potential checks include:
The review should begin with an appropriate pathway determination. A generic checklist is unlikely to reflect all device-specific requirements.
A useful finding should explain:
Supporting guide: AI Agent for FDA Submission Readiness: From Pathway Clarity to Gap Review
For EU medical devices, agents may help review alignment across areas such as:
The review should distinguish between regulations, guidance, standards and internal documentation.
A finding should show:
Supporting guide: AI Agent for MDR Compliance: Classification, Technical Documentation and Traceable Review
Regulatory agents can also support quality management system gap analysis.
A review may examine whether:
The system may identify two types of issue:
These findings require different corrective actions and should be reviewed in the context of the company’s processes and applicable requirements.
The terms used for regulatory technology often overlap. The clearest distinction is the type of work each approach supports.
| Approach | What it generally does well | Main limitation | Appropriate use |
| General-purpose AI assistant | Summarisation, rewriting and basic document support | May lack controlled regulatory sources and task-specific logic | Low-risk supporting work with human verification |
| Regulatory database or search tool | Provides access to records and source documents | Interpretation and workflow execution remain with the user | Expert-led research |
| Point regulatory AI tool | Automates one defined activity | May not connect decisions across the lifecycle | High-volume, repeatable specialist tasks |
| Rules-based regulatory software | Applies predefined decision trees and workflows consistently | Can be rigid and dependent on correct user interpretation | Stable, well-defined processes |
| AI regulatory affairs agent | Executes multi-step tasks using regulatory context, tools and sources | Requires governance and expert oversight | Complex research, drafting and review workflows |
| Regulatory consultant | Provides judgement, accountability and strategic interpretation | Capacity, cost and turnaround may be limiting | Novel, ambiguous or commercially strategic decisions |
A regulatory team may use several of these approaches together.
General AI can support editing. Databases can provide source access. Specialised agents can accelerate analysis. Consultants and internal experts can address ambiguity, strategic choices and accountability.
Teams should identify which parts of the workflow can be accelerated while preserving quality, traceability and regulatory control.
A credible AI regulatory affairs agent combines current and controlled regulatory knowledge, task-specific logic, traceable reasoning and evidence, consistency across connected documentation, appropriate human oversight, and clear limits on what the system can determine.
The system should retrieve relevant regulations, guidance, standards or database records rather than rely only on model memory.
Teams should ask:
The workflow should reflect the regulatory task being performed.
Classification, predicate analysis, clinical-evidence planning and quality-system review require different inputs and reasoning steps.
Teams should ask the provider to explain:
Users should be able to trace:
When the system flags a gap, it should indicate where the issue appears and which requirement supports the finding.
Regulatory documents should be reviewed in context.
A claim in one document may need to align with:
The agent should identify conflicts across the document set or state when its review is limited to one file.
Human approval is required at material decision points.
The system should not imply that:
The manufacturer remains responsible for the regulatory decisions and documentation.
A credible platform should identify what it cannot determine.
Examples include:
Clear limitations help teams decide when further expertise or regulator engagement is needed.
AI agents work best as analytical and productivity tools within a controlled regulatory process.
They do not replace:
Startups can use agents to structure regulatory questions earlier, explore plausible pathways and understand which requirements may affect development.
Useful applications include:
Internal teams can use agents to:
The benefit is both additional capacity and more systematic regulatory work. Complex, interconnected requirements can make gaps or inconsistencies difficult to spot until they create significant rework. AI agents can help teams identify these issues earlier and review information more consistently, while regulatory experts focus on resolving ambiguity, assessing risk and advising product teams.
Consultants can use agents to:
The consultant still provides strategic interpretation.
For example, a company may need to choose between a narrower initial claim that supports an earlier route to market and a broader claim that requires more evidence.
An agent can analyse the regulatory implications of each scenario. The consultant integrates that into a borader advice on which route best fits the company’s strategy, resources and risk tolerance.
The best time to begin is when the product concept is defined well enough to support meaningful regulatory analysis and before major development decisions become difficult to reverse.
An agent is particularly relevant when a team needs:
A specialised agent may be unnecessary for simple editing, summarisation or administrative work that does not require regulatory interpretation.
Guideways uses four specialised agents across connected stages of the medical device regulatory lifecycle.
Sherpa supports:
It uses the device description, intended use / purpose and indications for use to establish a structured regulatory direction.
Drafter uses device and regulatory information already available in the platform to create first drafts of technical and quality documents.
Examples include:
These outputs require review and approval by the responsible team.
Reviewer analyses technical file submissions, quality systems and deelopment documentation to identify:
Researcher answers questions using regulatory and project context.
It supports users in identifying and interpreting relevant evidence from regulatory sources and project documentation.
The agents share project and regulatory context.
Drafter can use information and regulatory analysis established through Sherpa. A clinical evaluation plan, for example, can begin with the device description, pathway analysis and identified requirements already recorded in the platform.
Researcher and Reviewer can also work with the regulatory sources and project documentation available within the same context.
This continuity can reduce contradictions between early strategy, document preparation and later review. The team remains responsible for confirming the information and approving the outputs.
Before selecting a platform, ask:
AI regulatory affairs agents can support research, pathway analysis, drafting and document review across the medical device lifecycle.
Their value depends on five factors:
Teams should assess whether the system produces work that regulatory professionals can inspect, challenge and approve.
Used within a controlled process, these agents increase regulatory capacity while keeping judgement and accountability with the people responsible for the device.
See how Guideways applies regulatory logic, source material and project context to classification, pathway planning, document drafting or submission review.